
Downloads · 30 days
16
9% of all-time downloads
Acellera/AceFF-1.1
AceFF-1.1 is a machine learning model from Acellera. Use it for the machine learning task on the model card, and read the license before you ship it in a product. The card lists the license as apache-2.0.
Organization(s): Acellera Therapeutics, inc Contact: [email protected] License: apache 2.0
Downloads · 30 days
16
9% of all-time downloads
All-time downloads
181
Public
Repo size
6.8 MB
Likes
4
Public
Click a slice to open those files.
.ckpt6.5 MB · 99%
From the Hugging Face model README
Organization(s): Acellera Therapeutics, inc
Contact: [email protected]
License: apache 2.0
Acellera AceFF 1.1 is a next-generation Neural Network Potential (NNP) designed for Relative Binding Free Energy (RBFE) calculations in drug discovery. It addresses key limitations of traditional molecular mechanics (MM) force fields and earlier NNP models, including restricted atom types, limited charge support, and computational inefficiencies.
The model leverages the TensorNet architecture[1] and the NNP software library TorchMD-Net [2] to provide accurate predictions for diverse drug-like compounds, supporting all key chemical elements and charged molecules. AceFF 1.1 improves the stability of molecular dynamics simulations, supports 2 fs timesteps, and achieves state-of-the-art accuracy with fewer outliers in RBFE predictions.
AceFF 1.1 is the second version of a new family of potentials released by Acellera. It uses TensorNet 1-layer trained on Acellera's internal proprietary dataset of molecular forces and energies using the wB97M-V/def2-tzvppd level of theory and VV10 dispersion corrections.
The training set was built on PubChem. We extracted the SMILES and generated molecules, filtering out molecules larger than 30 atoms. We kept only molecules with the elements H, B, C, N, O, F, Si, P, S, Cl, Br, and I.
The AceFF1.1 dataset includes an additional 1 million conformations of molecules with up to 30 atoms and more diverse charges, building upon the AceFF1.0 dataset.
The table shows the results on the Wiggle150 benchmark. We include AIMNet2 and ANI-2x for comparison.
| Method | MAE (kcal/mol) | RMSE (kcal/mol) |
|---|---|---|
| AceFF1.1 | 2.51 | 3.18 |
| AceFF1.0 | 2.73 | 3.32 |
| AIMNet2 | 2.39 | 3.13 |
| ANI-2X | 4.41 | 5.41 |
Performance of NNPs on Wiggle150 benchmark
We create our own hold-out test set by labelling 650 ligands from the Schrodinger public binding free energy benchmark (Jacs, Merk, and charge_annhil sets) with AceFF DFT level of theory. We evaluate the Force MAE of the AceFF predictions.
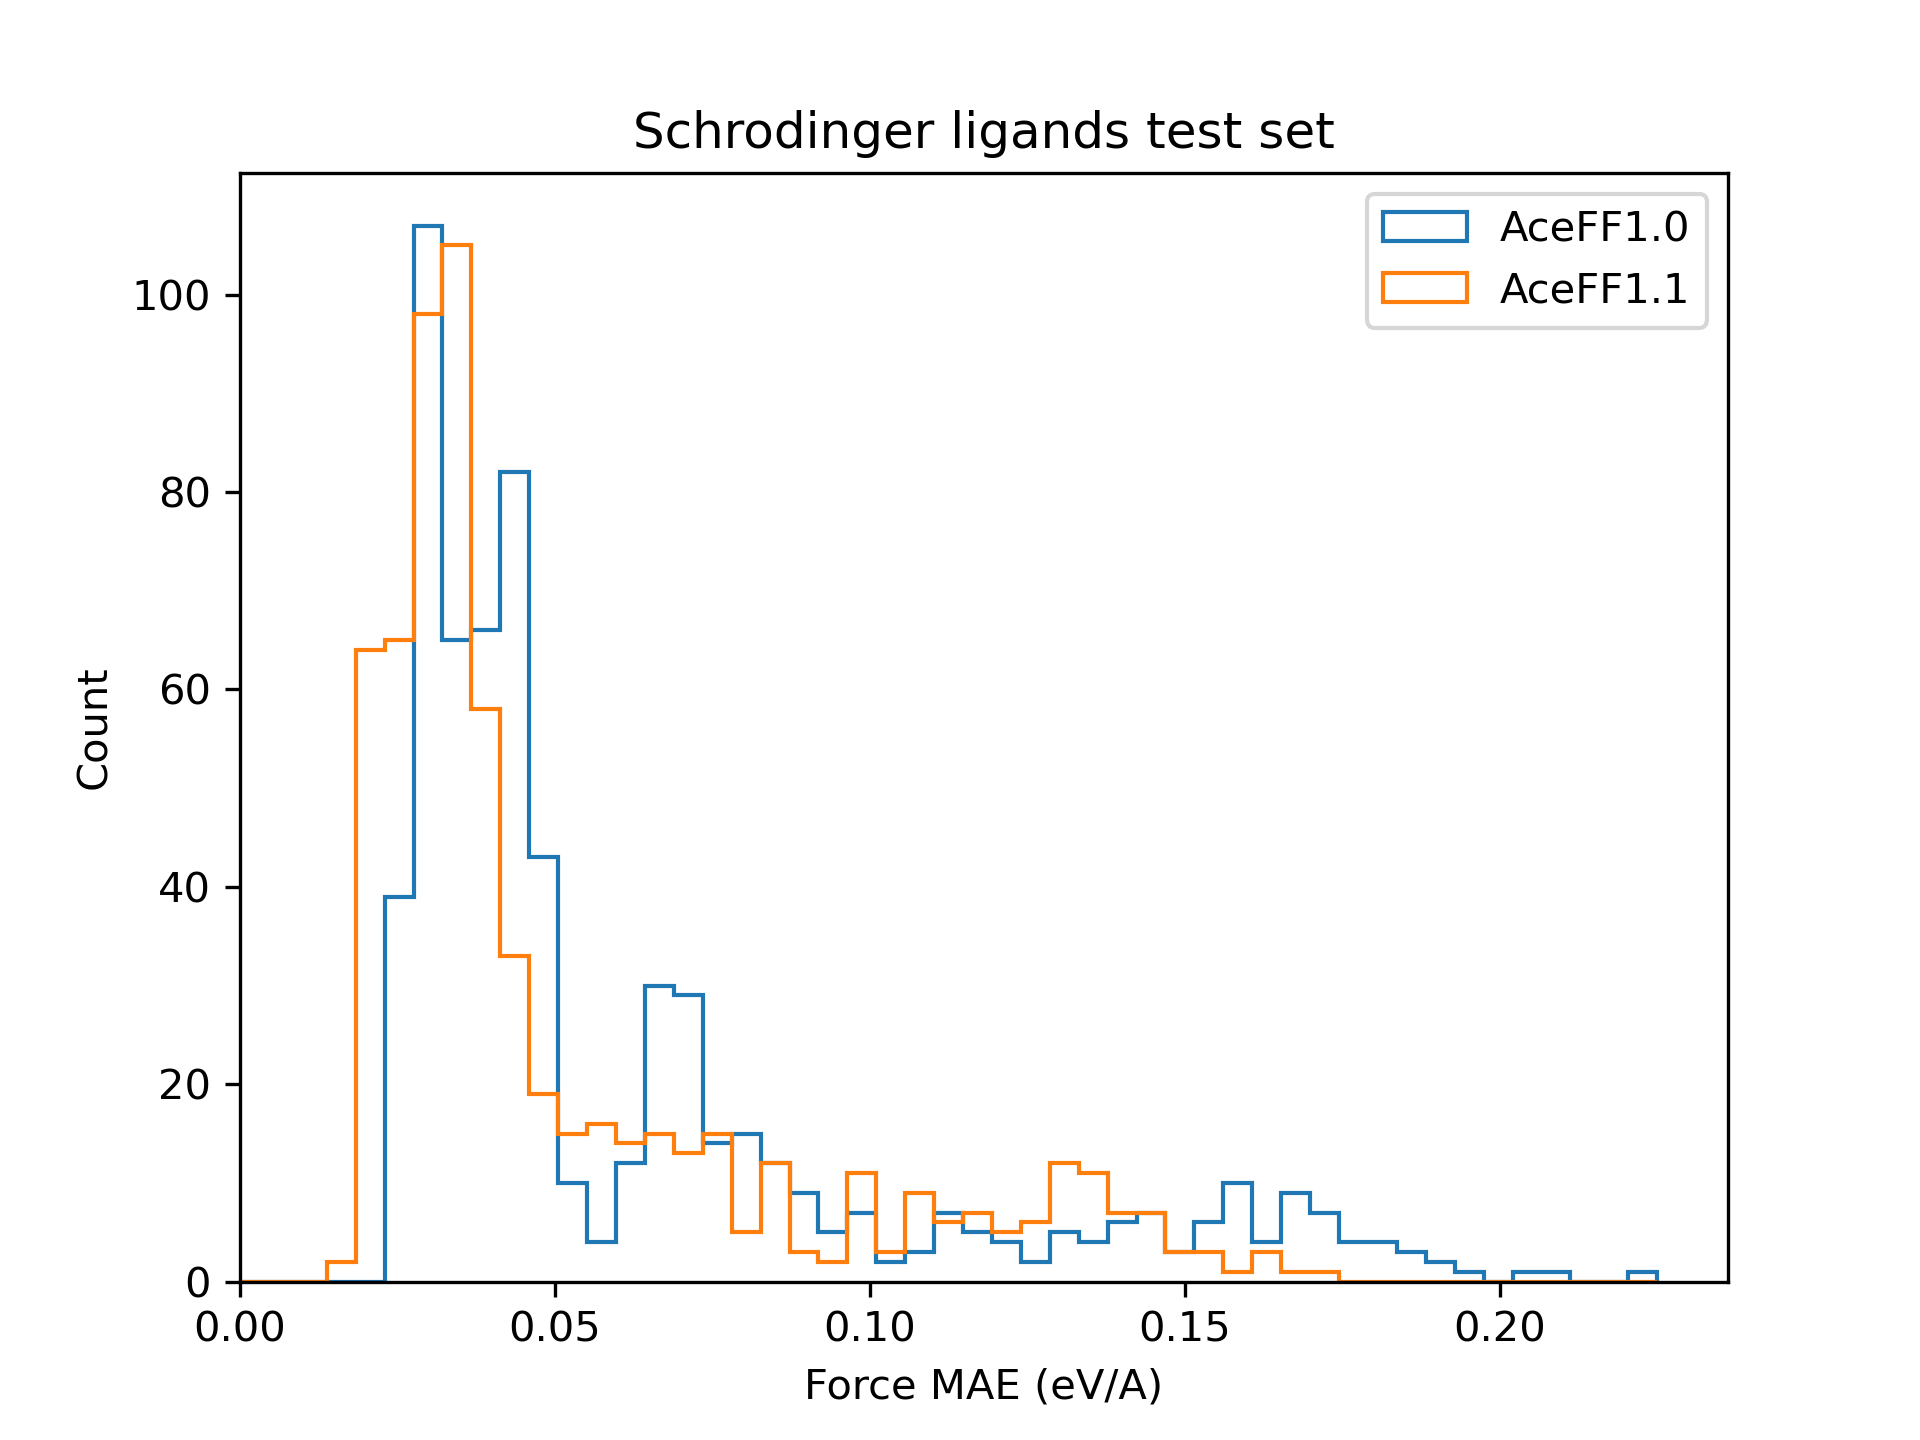
AceFF 1.1 is designed for use alone or in an NNP/MM approach, where the ligand is treated with the neural network potential and the environment with molecular mechanics.
Example notebooks are available in Google Colab, demonstrating the use of AceFF with OpenMM and ASE.

Run ML potential molecular simulations of a small molecule using ACEMD with this tutorial , e.g. to minimize.
For a tutorial on running mixed protein-ligand simulations, refer to NNP/MM in ACEMD.
[1] Simeon, Guillem, and Gianni De Fabritiis, Tensornet: Cartesian tensor representations for efficient learning of molecular potentials, Advances in Neural Information Processing Systems 36 (2024), https://arxiv.org/abs/2306.06482
[2] Raul P. Pelaez, Guillem Simeon, Raimondas Galvelis, Antonio Mirarchi, Peter Eastman, Stefan Doerr, Philipp Thölke, Thomas E. Markland, Gianni De Fabritiis, TorchMD-Net 2.0: Fast Neural Network Potentials for Molecular Simulations, J. Chem. Theory Comput. 2024, 20, 10, 4076–4087, https://arxiv.org/abs/2402.17660
[3] Francesc Sabanés Zariquiey, Stephen E. Farr, Stefan Doerr, Gianni De Fabritiis, QuantumBind-RBFE: Accurate Relative Binding Free Energy Calculations Using Neural Network Potentials, https://arxiv.org/abs/2501.01811 (2025).